Windchill PLM Software for Medical Device Manufacturers: Compliance Features Explained
How Windchill PLM software supports medical device teams under FDA's QMSR: design control, change, CAPA, complaints and e-signatures, plus its limits.
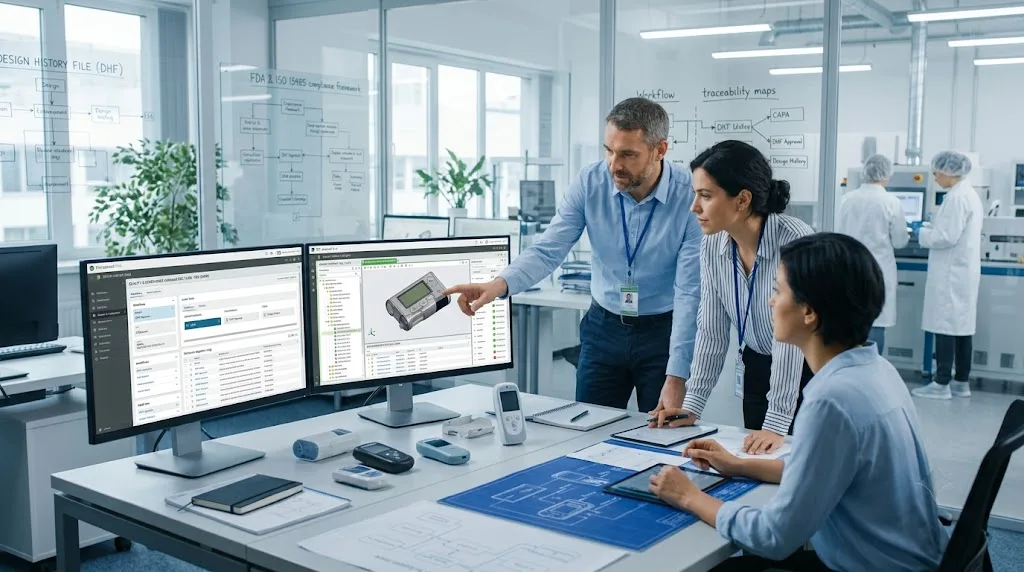
Windchill PLM software helps medical device manufacturers control the records FDA inspectors ask for: design history, change approvals, risk links, complaints and corrective actions. It does not make a company compliant by itself. This article maps Windchill's medical device features to FDA and ISO 13485 requirements and marks where your own procedures still carry the load.
Since February 2, 2026, the Quality Management System Regulation (QMSR) has made ISO 13485:2016 the backbone of 21 CFR Part 820, as FDA describes it. The sections below therefore follow ISO 13485 clauses instead of the old QSR subparts.
What the QMSR Changes for a Medical Device PLM System
The QMSR changes structure and vocabulary more than substance, but it widens what a PLM system must be ready to show. FDA kept added provisions on top of ISO 13485, including unique device identification (UDI), traceability, medical device reporting and corrections and removals. It also dropped the old exception that kept internal audit and management review reports away from inspectors, according to a Covington analysis. Audit findings now belong in a system that can produce them cleanly.
Terminology is the quieter trap. Teams still say design history file (DHF) and device master record (DMR), while ISO 13485 organizes the same material as design and development files and a medical device file [VERIFY]. PTC's documentation still uses the DHF and DMR names, so expect to map each name to a clause.
Windchill Quality Solutions Mapped to ISO 13485
PTC groups its medical device functions into three module sets: document control, design control, and surveillance with corrective action, according to PTC's help center. The table pairs each capability with the requirement it serves and a question worth asking in a demo.
Because Windchill sits on the product structure, a complaint can be tied to a BOM revision. A quality engineer then sees which design and change record produced the affected unit. That link is the practical case for medical device PLM over a standalone quality tool.
Electronic Signatures: What the Software Can and Cannot Do
Windchill can supply technical controls for 21 CFR Part 11, but Part 11 attaches to your records, procedures and validation, not to a product. FDA's 2003 scope and application guidance says the agency interprets the rule narrowly, focusing on records kept electronically under other FDA requirements. The design control module includes electronic signatures, yet signature meaning, role design and user training remain your documents. FDA compliance rests on validating the configured system, not the vendor's brochure.
Where Compliance Projects Go Wrong
These failure modes repeat often enough to plan for (analysis, not survey data):
Configuration drift. Heavy workflow customization expands validation scope at every upgrade. Stay close to standard modules wherever a procedure allows.
Split records. BOMs in PLM and CAPA in a separate system break traceability the moment someone retypes a part number.
Automating a weak process. A vague change procedure becomes a vague workflow, only faster.
Choosing a Starting Point for Windchill PLM Software
The right first module depends on where your quality system stands today (analysis, hypothetical situations):
One device, early stage: begin with document control, design control and change management. Add post-market modules before shipping.
Existing eQMS: run a one-question test. From a single complaint, can you reach the BOM revision, change record and CAPA without leaving the system? If not, cost out consolidation.
Multi-site or contract manufacturing: prioritize controlled supplier access and UDI data before advanced features.
Sequence matters because configuration drives validation effort. Map current records to ISO 13485 clauses first, validate document and change control second, then add design control, then complaints and CAPA. Teams without in-house PTC expertise can scope the configuration with an implementation partner before buying; 3HTi's Windchill software page shows what that scope can include.
Conclusion
Windchill PLM software gives device teams the structure to produce inspection-ready records, and the QMSR raises the stakes on having them. Compliance still lives in procedures, validation and training. Pick one recent complaint and trace it through your current systems to the design, change and corrective action behind it. Wherever the trail breaks is your roadmap.
0 comments
Log in to leave a comment.
Be the first to comment.